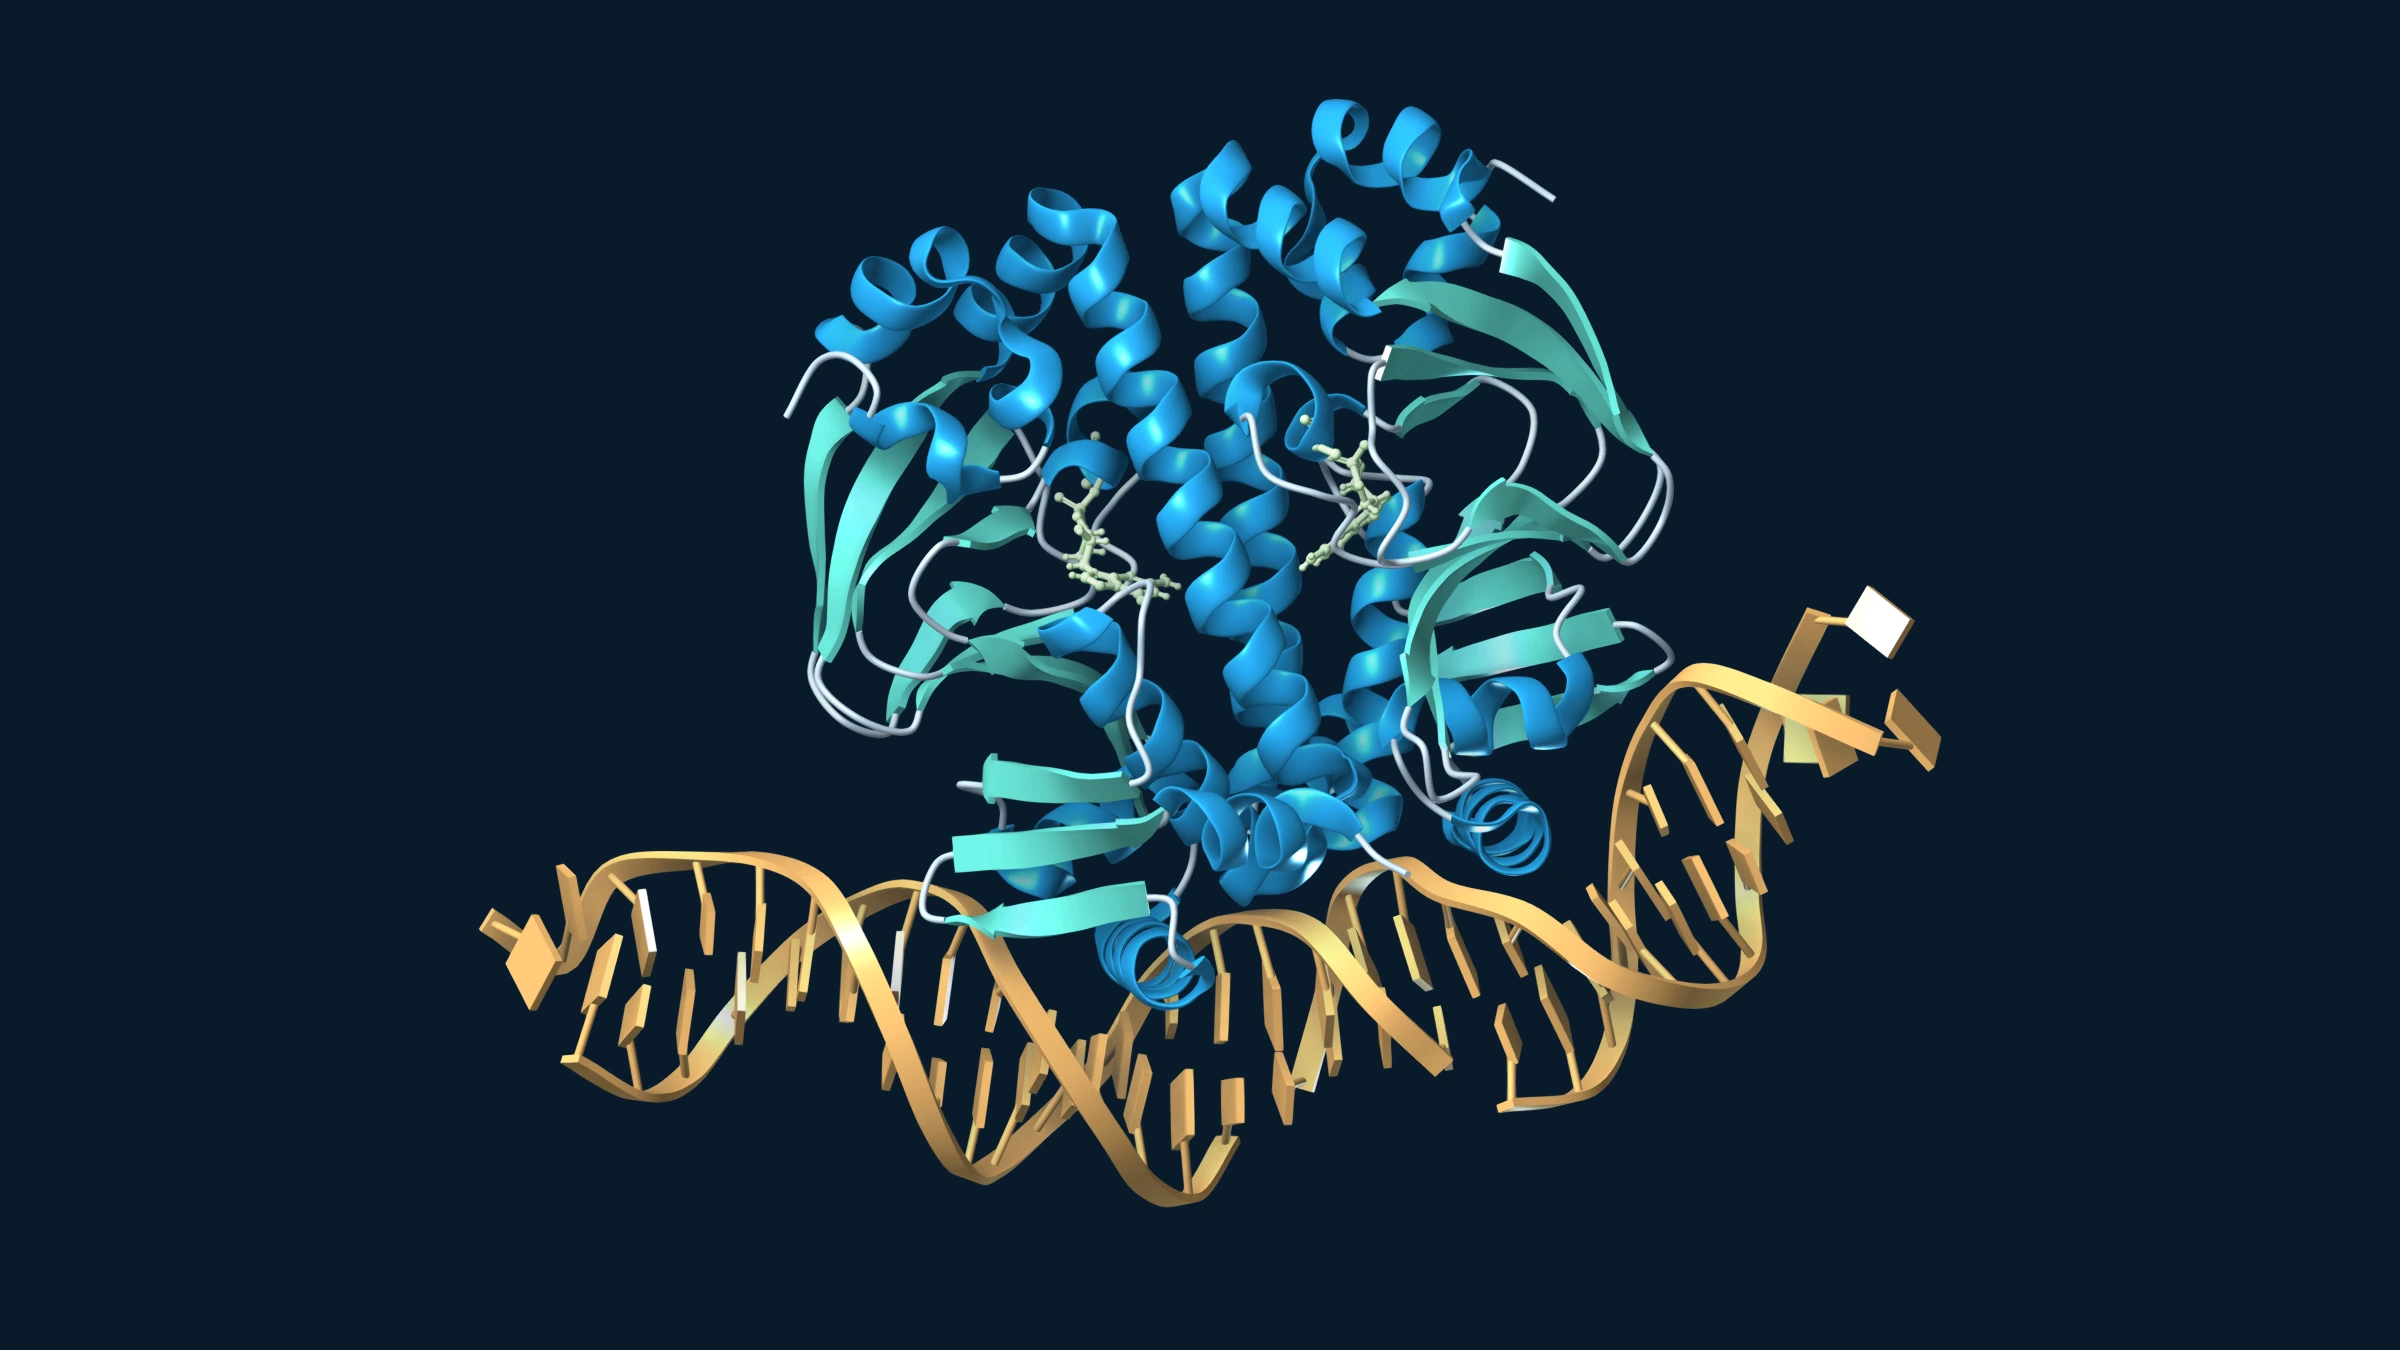
How do proteins fit together with DNA, RNA and smaller molecules? AlphaFold 3, introduced by Google DeepMind and Isomorphic Labs in May 2024, extends structure prediction to these mixed molecular complexes. The aim is to give researchers testable models of how the components might be arranged in three dimensions. Launch announcement.
What AlphaFold 3 adds
The study in Nature describes a model that predicts complexes containing proteins, nucleic acids, small molecules, ions and modified residues. Earlier AlphaFold-based methods, including AlphaFold-Multimer, could already predict protein complexes. The important expansion is the range of molecular components that can be modelled together.
On the authors’ benchmarks, AlphaFold 3 improved performance over the selected comparison methods for several interaction types, including protein–ligand and protein–nucleic acid complexes. These are results for defined tests; they do not mean every prediction will be correct. Abramson and colleagues, 2024
From molecular inputs to a proposed structure
AlphaFold 3 uses a diffusion-based process to generate atomic coordinates. Starting from a noisy arrangement, it repeatedly refines positions using information about the supplied molecules. The resulting model offers a proposed arrangement of the complex. This computational sequence should not be read as a movie of how the molecules actually fold or bind. Google DeepMind and Isomorphic Labs.
What researchers still need to check
The paper reports limitations including incorrect stereochemistry, clashes between atoms and overly ordered predictions for some disordered regions. It also cautions that the model does not describe molecular dynamics and tends to favour particular conformations. A plausible-looking structure can therefore omit behaviour that matters biologically. Abramson and colleagues, 2024.
Confidence measures help identify uncertain parts of a prediction. Local confidence, reported as pLDDT, addresses a different question from confidence in the relative placement of molecular partners. For an interface, predicted aligned error (PAE) and interface predicted TM-score (ipTM) provide more relevant checks. These estimates should guide scrutiny of the particular interaction being investigated. Official output documentation.
Why this matters for enzyme research
For an enzyme researcher, a proposed complex can help frame a specific experiment: which residues might contact a substrate, and which changes would test that interpretation? The value lies in turning a structural hypothesis into an experiment with a clear comparator.
Our practical interpretation is that a binding pose alone cannot establish catalytic activity. An enzyme variant still needs appropriate measurements of turnover, substrate preference and stability. Likewise, a predicted drug–target complex cannot by itself demonstrate treatment efficacy or safety. Structure prediction can help decide what to test, while experiments determine whether the proposed function is supported.
Access: a dated update
At launch in May 2024, AlphaFold Server offered a browser-based route to predictions for non-commercial research. Google subsequently announced the release of model code and weights on 11 November 2024, after this article’s original publication. Google’s dated announcement.
As checked on 20 September 2026, the official repository provides an AlphaFold 3 inference pipeline under Apache 2.0. Model parameters and outputs have separate terms, including non-commercial restrictions. The server remains another access route with its own conditions and a more limited selection of ligands and modifications. The code licence should therefore not be treated as permission for every use of the model or its outputs. Official repository, model-parameter terms.
Related reading: New Study Compares Four Enzymes for Plastic Breakdown.
Update note, 20 September 2026: This article was revised to qualify prediction-accuracy claims, clarify the earlier ability to model protein complexes, and distinguish launch-time access from the subsequent code release.
References
- Abramson, J., Adler, J., Dunger, J., et al. (2024). Accurate structure prediction of biomolecular interactions with AlphaFold 3. Nature, 630, 493–500. DOI: 10.1038/s41586-024-07487-w.
- Google DeepMind AlphaFold team and Isomorphic Labs (2024). AlphaFold 3 predicts the structure and interactions of all of life’s molecules. Published 8 May 2024; code-release update 11 November 2024. Announcement.
- Google DeepMind. AlphaFold 3 inference pipeline, output documentation and model-parameter terms. Checked 20 September 2026.
Image sources
- Werel, L., Farmani, N., Krol, E., Serrania, J., Essen, L. O., and Becker, A. (2023). Structural basis of dual specificity of Sinorhizobium meliloti Clr, a cAMP and cGMP receptor protein. mBio, 14, e03028-22. DOI: 10.1128/mbio.03028-22. Experimental coordinates: RCSB PDB 7PZB.
- Sehnal, D., et al. (2021). Mol* Viewer: modern web app for 3D visualization and analysis of large biomolecular structures. Nucleic Acids Research, 49, W431–W437. DOI: 10.1093/nar/gkab314. Visualization software used for the accompanying illustration.